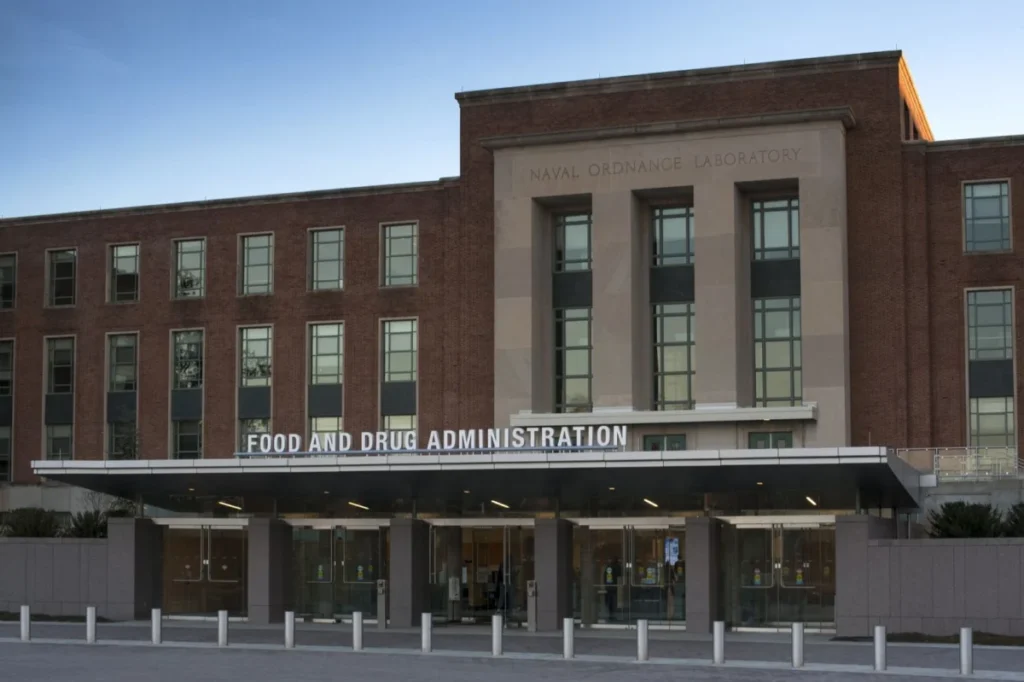
The U.S. Food and Drug Administration (FDA) has taken a significant step in clarifying the regulatory landscape for dietary supplements by finalizing a crucial portion of its draft guidance on New Dietary Ingredient Notifications (NDINs). This latest development addresses the required elements for submitting an NDIN, a process designed to ensure the safety of novel ingredients introduced into the supplement market. However, despite this progress, a central and long-standing question regarding the definition of a "new" dietary ingredient, particularly concerning chemical alterations, remains unresolved, leaving industry stakeholders in a state of continued uncertainty.
A Protracted Regulatory Journey
The journey toward updated NDIN guidance has been a complex and protracted one, marked by industry pushback, regulatory revisions, and a piecemeal approach to finalization. The initial draft guidance on NDINs was first released by the FDA in 2010. This inaugural document immediately ignited significant controversy within the dietary supplement industry and drew sharp criticism from members of Congress. The concerns were multifaceted, encompassing several key sticking points.
One prominent issue revolved around the classification of "synthetic botanicals," a term that has since been superseded by the more precise designation of "nature-identical" ingredients. Industry stakeholders questioned how such ingredients, which may be synthesized but chemically identical to naturally occurring compounds, should be regulated. Another significant point of contention was the requirement for separate NDINs for every new product containing a New Dietary Ingredient (NDI). This raised concerns about redundant submissions and potential burdens on manufacturers. Furthermore, debates ensued regarding whether the notification process should be exclusively ingredient-focused or also consider the finished product formulation.
The FDA’s response to this initial wave of feedback was to withdraw the 2010 guidance, acknowledging the need for a more thorough re-evaluation of the proposed framework. This withdrawal signaled a recognition of the significant concerns raised by the industry and the legislative branch.
A revised version of the draft guidance emerged in 2016. While this iteration aimed to address some of the previously identified issues, it did not achieve immediate universal approval from the industry. The subsequent discussions and engagements between the FDA and industry representatives led to a strategic shift in the agency’s approach. Rather than attempting to forge a single, comprehensive compromise that would satisfy all parties simultaneously, the FDA announced its intention to finalize the guidance in stages, addressing specific sections of the document incrementally. This "piece-by-piece" strategy was designed to allow for more focused deliberation and potentially faster progress on individual components of the complex regulatory framework.
Finalized Section VI: Detailing Notification Requirements
The announcement this week marks the official finalization of Section VI of the draft guidance. This section is dedicated to elaborating on the essential elements that must be included in a New Dietary Ingredient Notification (NDIN). For context, the original text within the Dietary Supplement Health and Education Act (DSHEA) of 1994 that pertains to NDIs is remarkably concise, comprising only 381 words. DSHEA, in fact, is often cited for its brevity in comparison to many other federal regulations. In stark contrast, the finalized guidance explaining this foundational legal provision now extends to a comprehensive 53 pages, illustrating the depth of detail the FDA intends to provide.
Section VI meticulously outlines the information that manufacturers and distributors must submit to the FDA when notifying the agency of a new dietary ingredient. This includes detailed specifications on how the identity of an ingredient should be established, ensuring clarity and consistency in its characterization. The guidance also mandates a significant level of detail regarding the manufacturing process. This is crucial for understanding how the ingredient is produced, potential impurities, and its overall quality. Furthermore, the finalized section provides specific information tailored to various categories of ingredients, including enzymes, botanicals, extracts, salts, and microbial ingredients, recognizing the unique considerations associated with each.
Following the detailed instructions on establishing ingredient identity, the guidance dedicates substantial attention to the methodologies for demonstrating the safety of these new ingredients. This critical aspect of the NDIN process includes in-depth discussions on the significance of "history of use" as a form of safety evidence, defining what constitutes "other evidence of safety" beyond historical use, and addressing various other relevant scientific and toxicological considerations. The aim is to equip submitters with a clear understanding of the types of data and scientific rigor expected to assure the FDA that a new dietary ingredient can be safely consumed.
The Lingering Shadow of "Chemical Alteration"
Despite the significant progress represented by the finalization of Section VI, the document conspicuously avoids addressing a particularly contentious and persistent ambiguity: the threshold for "chemical alteration" that would necessitate the filing of a new NDIN. This remains a central question that has plagued the industry since the initial guidance was proposed.
The core of the issue lies in defining when a modification to an existing ingredient, particularly through different extraction or processing methods, renders it sufficiently "new" to require a fresh safety assessment and notification to the FDA. For instance, if a company transitions from a simple water extraction process to one employing solvents, the resulting extract might be chemically altered. The critical question is whether this alteration is substantial enough that the original safety data, established for the water-extracted form, may no longer be applicable. Similarly, the use of more advanced techniques like supercritical fluid extraction raises similar concerns about potential chemical modifications and their regulatory implications.
This ambiguity was a major source of apprehension during the initial development of the NDIN guidance in 2010. At that time, there was a significant fear within the industry that an overly broad interpretation by the FDA of what constitutes a "new" ingredient due to chemical changes could lead to a cascade of new NDIN filings. Such a scenario could potentially render thousands of products already legally on the market as "adulterated" by definition, creating widespread disruption and legal challenges. The lack of clear parameters for chemical alteration has therefore been a significant hurdle for innovation and product development, forcing companies to navigate a landscape of uncertainty.
Implications for the Supplement Industry
The finalized Section VI provides much-needed clarity on the procedural and evidential requirements for NDINs. Manufacturers and formulators can now proceed with greater confidence in understanding the specific information the FDA expects regarding ingredient identity, manufacturing processes, and safety substantiation. This clarity is expected to streamline the submission process for genuinely novel ingredients and enhance the overall quality of NDINs submitted to the agency. The detailed guidance on various ingredient types also suggests a more nuanced and scientifically informed approach by the FDA.
However, the continued absence of definitive guidance on chemical alteration means that a significant regulatory gray area persists. This uncertainty may continue to stifle innovation, as companies may be hesitant to invest in research and development involving modified extraction or processing techniques without a clear understanding of the regulatory implications. The potential for future enforcement actions based on differing interpretations of what constitutes a "new" ingredient due to chemical changes remains a concern.
The FDA’s decision to proceed with a piecemeal finalization strategy, while allowing for progress on specific aspects of the guidance, also means that the overall regulatory framework for NDIs is still incomplete. The industry will be eagerly awaiting further updates and finalizations of other sections of the draft guidance, particularly those that address the remaining contentious issues. The hope is that future iterations will provide the comprehensive clarity necessary for a robust and predictable regulatory environment that fosters both consumer safety and industry growth. This ongoing regulatory dialogue underscores the complex balance the FDA must strike between ensuring public health and supporting a dynamic and evolving marketplace for dietary supplements. The final resolution of the chemical alteration question will be a critical determinant in shaping the future of ingredient innovation and compliance within the U.S. dietary supplement industry.